materials-chemistry/ai in a supporting role/Chemistry of Materials 2022 · v2
Machine learning assigns atomic charges to size up polarisation in bismuth vanadate conductors
Researchers probed the local atomic arrangement of two tin- and germanium-substituted bismuth vanadate oxide ion conductors. A Gaussian process model, trained on charges from quantum calculations, assigned charges across atomic models too large for those calculations to handle.
spectrum · one line per step, placed by what the step does · bright lines used AI
Local Structure in α-BIMEVOXes (ME = Ge, Sn)
Chemistry of Materials, 2022
doi:10.1021/acs.chemmater.2c03001 · record aix-00159 v2 · checked 2026-10-09
- AI was for
- Property prediction, Simulation surrogate
- Model family
- Gaussian process
- Checked by
- Held-out
- Code
- not reported
AI processed or interpreted data, but the main finding does not rest on it.
What this research was about
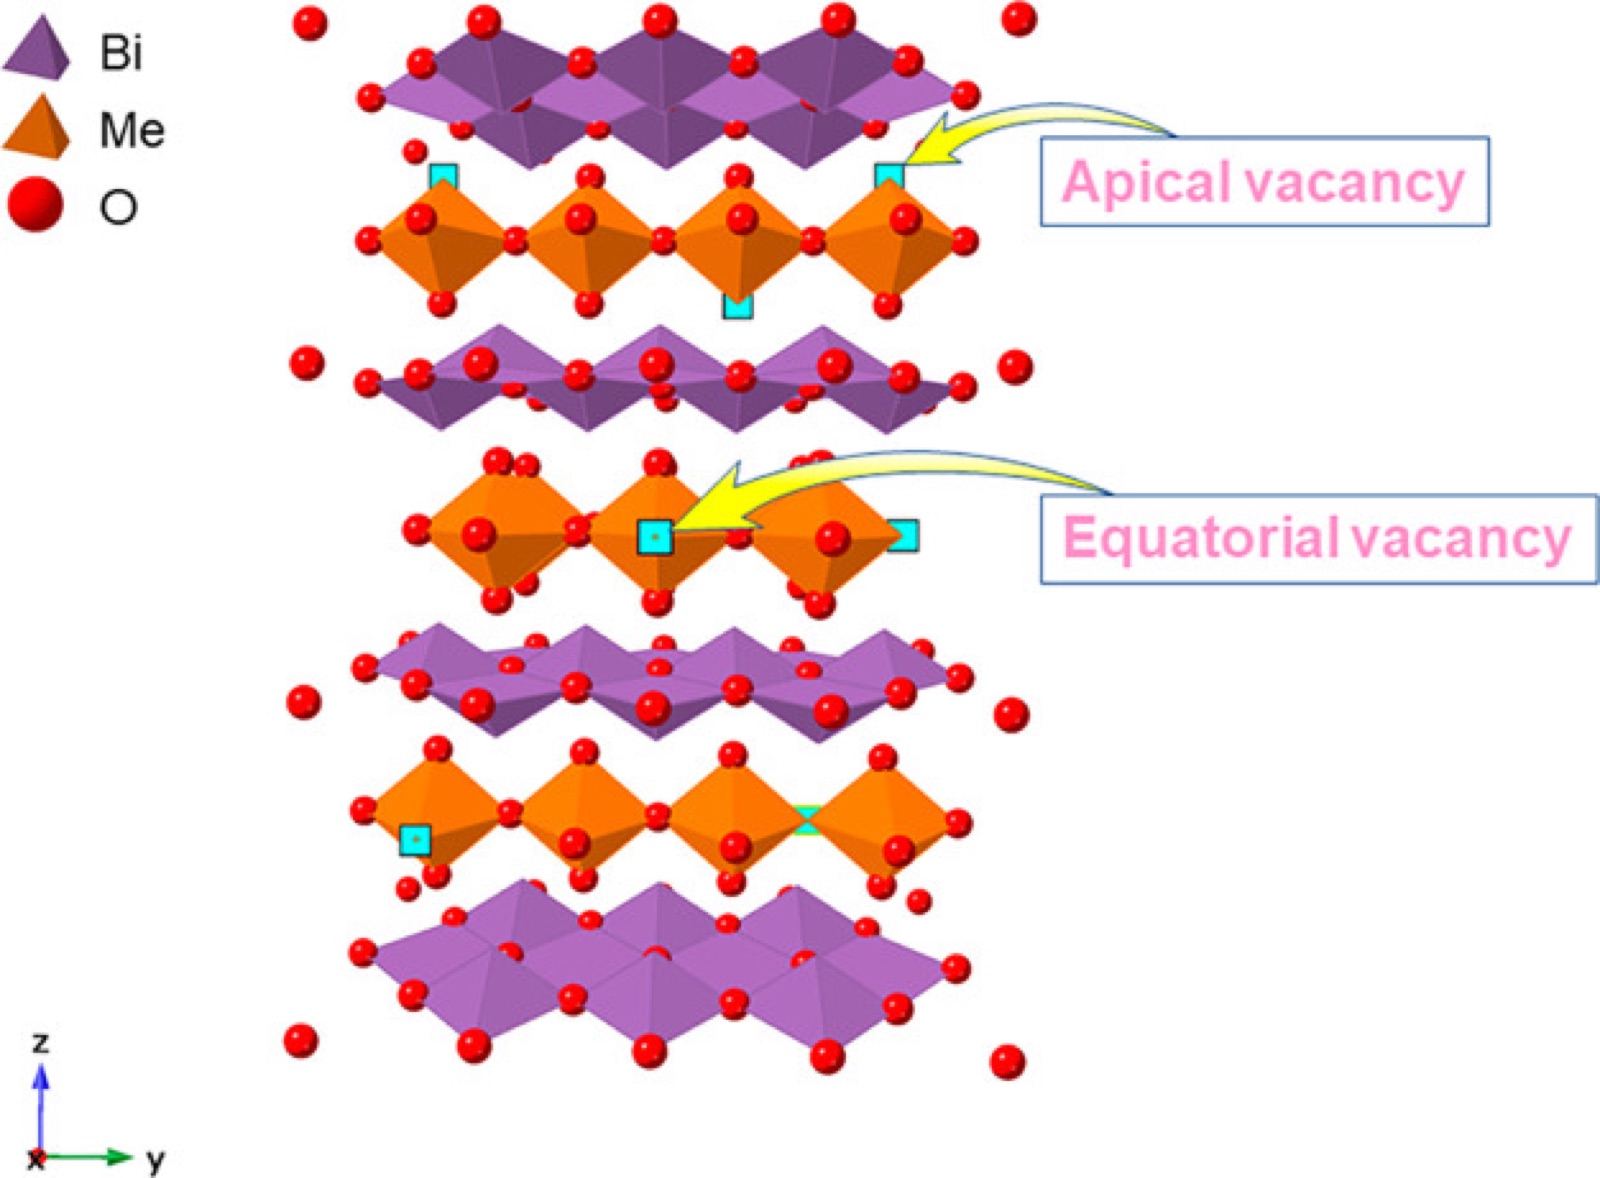
Some ceramics conduct electricity not by moving electrons but by shuffling oxygen ions through gaps in their crystal structure. The BIMEVOX family, based on bismuth vanadium oxide with other elements substituted in, is one such group. What makes these materials work is partly disorder: oxygen sites are left empty, and the atoms around them sit at angles and distances that vary from place to place. An average picture from standard crystallography smooths all that away. To see the local arrangement, researchers combine scattering experiments with computer models of thousands of atoms that are adjusted until they match the measured data.
The team studied two compositions, one substituted with germanium and one with tin. They collected neutron and X-ray total scattering data at room temperature and at 700 °C, built atomic models by reverse Monte Carlo fitting, and added solid-state nuclear magnetic resonance and impedance measurements. They also asked whether the structure is polar, meaning the positive and negative charges inside it are offset rather than balanced, and measured dielectric permittivity alongside.
Where AI came in
Working out whether a structure is polar means knowing how much charge each atom carries and how far it has shifted from a symmetric reference position. Charges can be extracted from quantum mechanical calculations, but only for small groups of atoms; the reverse Monte Carlo models here contained about 10 000 atoms, beyond what those calculations could treat. So the researchers ran density functional calculations on small configurations, extracted partial ionic charges, and described each atom's surroundings with a numerical fingerprint called a SOAP descriptor.
A Gaussian process regression model, a statistical method that learns a smooth relationship between inputs and outputs, was fitted to those descriptor and charge pairs using Scikit-learn, with the data split in a 0.8 to 0.2 ratio between training and validation. The fitted model then predicted a charge for every atom in the large configurations. Those charges, combined with how far each atom sat from an ideal symmetric reference, gave spontaneous polarisation values along each lattice direction, all below 2 μC cm per unit area. The model stood in for the quantum calculation, not for the structural analysis itself.
Written by AIxSci from the checked record below, to give context for readers outside the field. It is not part of the record.
The work
Technical · from the record
Two tetravalent-substituted BIMEVOX oxide ion conductors, Bi2V0.9Ge0.1O5.45 and Bi2V0.95Sn0.05O5.475, were characterised by neutron and X-ray total scattering with reverse Monte Carlo modelling, solid-state NMR and impedance spectroscopy. Because the reverse Monte Carlo configurations of about 10 000 atoms were too large for ab initio treatment, a Gaussian process regression model was fitted to Bader partial charges from density functional calculations and used to assign charges across those configurations, from which spontaneous polarization was computed and compared with ideal centrosymmetric models. The calculated polarization values were below 2 μC cm per unit area along the x, y and z directions, and dielectric permittivity measurements were interpreted as indicating weakly ferroelectric character for the α-phase. The reverse Monte Carlo models indicate Ge adopts tetrahedral geometry at 25 and 700 °C, while Sn is mainly six-coordinate at 25 °C and four-coordinate at 700 °C.
How AI was used
Density functional calculations in VASP (PBE molecular dynamics at 1500 K, structural relaxation, then PBE0 single-point calculations) were used to obtain Bader partial ionic charges for small configurations of each composition. Training and validation sets were built from those charge values together with the atomic environment of each atom encoded with the Smooth Overlap of Atomic Positions descriptor, split in a 0.8:0.2 ratio. A Gaussian process regression model with a radial basis function kernel, implemented in Scikit-learn, was fitted to this data and then applied to the much larger reverse Monte Carlo configurations, which exceed the size tractable by the ab initio method. The predicted per-atom charges, combined with atomic displacements relative to ideal centrosymmetric reference structures after folding the configurations back onto the crystallographic unit cell, were used to compute spontaneous polarization along each lattice direction. No learned model was involved in the reverse Monte Carlo fitting, Rietveld refinement, NMR fitting or impedance analysis.
The shape of the work
Structural · the record, drawn
no AI
Collect neutron and X-ray total scattering data
Obtaining raw data, whether by measurement, download or retrieval.
data collections corresponding to proton beam charges of ca. 1000 μA h were made to allow for total scattering analysiswhere the paper describes this · verbatim
no AI
Reverse Monte Carlo modelling of local structure
Numerical or physics simulation, including where a learned surrogate replaces it.
The reverse Monte Carlo (RMC) method using the RMCprofile software was applied to model local structure.where the paper describes this · verbatim
no AI
Density functional calculations and Bader charge analysis
Numerical or physics simulation, including where a learned surrogate replaces it.
The calculation of ionic charges was performed using the Bader partitioning scheme.where the paper describes this · verbatim
no AI
Encode atomic environments and build training and validation sets
Encoding data into features, descriptors, embeddings or graphs.
the atomic environments of each atom, encoded using the Smooth Overlap of Atomic Positions descriptorwhere the paper describes this · verbatim
AI
Train Gaussian process regression charge model
Fitting model parameters, including fine-tuning an existing model. The AI stood in for simulation.
The machine learning model was created based on the Gaussian Process Regression approach, with the radial basis function kernelwhere the paper describes this · verbatim
AI
Predict charges across large RMC configurations
Running a trained model over new data to predict, classify or score. The AI stood in for simulation.
The RMC structures were then used for the prediction of charges using a machine learning method as described in the Experimental Section.where the paper describes this · verbatim
no AI
Calculate spontaneous polarization from predicted charges
Extracting understanding from model behaviour.
compared with the ideal centrosymmetric structural models to obtain the spontaneous polarization, Ps, value along each lattice directionwhere the paper describes this · verbatim
What the record says
Technical · every part carries its own basis
+ in the paper~ our reading− not reported
How to read the quotations. A quotation shows where the paper describes something. It does not quote every value beside it: one passage locates a part of the work, and values without their own quotation are our reading of that passage.
A Gaussian process model supplied partial ionic charges used to compute spontaneous polarization; the paper's structural conclusions rest on diffraction, RMC and NMR, and the polar character is also supported by dielectric measurements
The machine learning model was created based on the Gaussian Process Regression approach, with the radial basis function kernel as implemented in Scikit-learn.where the paper describes this · verbatim
Training and validation sets were weighted in a 0.8:0.2 ratio.where the paper describes this · verbatim
Neutron data used in this work are available at https://doi.org/10.5286/ISIS.E.RB1820126.where the paper describes this · verbatim
What this paper did not report
Technical · absence is published deliberately
- CodeWhether the code is available is not stated.
- Trained model weightsWhether the trained model is available is not stated.
- ComputeThe hardware or time used is not stated.
- How many were testedThe paper gives no count of what was tested.
- Version of Gaussian process regression model for ionic charges (SOAP descriptors, RBF kernel, Scikit-learn)Which version of the model was used is not stated.
About this article
Record aix-00159, version 2, checked by a person on 2026-10-09. The record describes the paper; it does not assess whether the paper's findings are right. The paper is published under CC-BY; quotations are at most 25 words. How we work · Report an error